
🟢 Strong Evidence
Mitochondria have long been understood as the cell’s power plants, responsible primarily for generating adenosine triphosphate (ATP), the universal energy currency. However, emerging evidence from cellular and molecular biology research reveals that mitochondrial function extends far beyond energy production—these organelles act as central hubs coordinating cell death, immune activation, metabolic sensing, and epigenetic gene regulation. This expanded understanding reframes mitochondrial dysfunction not merely as an energy crisis, but as a failure of multiple cellular communication systems simultaneously.
Key takeaways
- Mitochondria release cytochrome c to trigger programmed cell death (apoptosis) when cells are irreparably damaged
- Mitochondrial redox signaling via hydrogen peroxide and NAD+/NADH ratios regulates protein modification and cellular stress responses
- Mitochondrial DNA and metabolites activate innate immune pathways and control chromatin remodeling for cell-fate decisions
- Dysfunction in any of these signaling axes—not energy production alone—may drive disease pathology
Mitochondrial Functions Beyond Energy Production
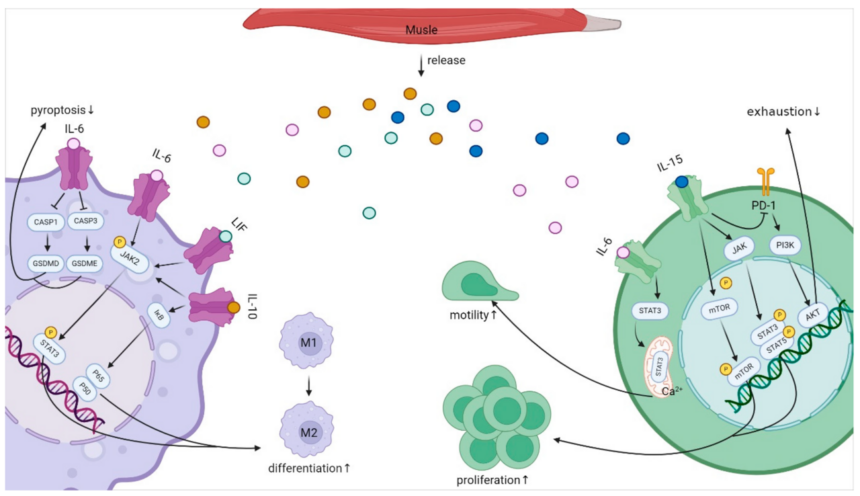
Five parallel signaling axes controlling cell survival, immunity, and gene expression
Illustration based on cellular biology consensus | Georgian Medical Journal News

Apoptosis and Cytochrome c: The Death Switch
When a cell suffers irreparable DNA damage or receives external death signals, mitochondria respond by releasing the protein cytochrome c into the cytoplasm. This molecule acts as a molecular alarm, binding to apoptosis protease-activating factor 1 (Apaf-1) and triggering the caspase cascade—a proteolytic pathway that systematically dismantles the cell in a controlled manner. Research in cell biology literature demonstrates that this mitochondrial gate-keeping function is essential for preventing the survival of transformed or damaged cells that might otherwise become cancerous. Without functional cytochrome c release, cells escape death signals and can accumulate mutations unchecked.
The balance between pro-survival and pro-death Bcl-2 family proteins determines whether cytochrome c reaches the cytoplasm. This decision point resides entirely within the mitochondrial outer membrane, making mitochondria arbiters of whether a cell lives or dies—a function entirely separable from energy production.
Redox Signaling and Metabolic Thiol Switches
Mitochondria generate reactive oxygen species (ROS) including hydrogen peroxide (H₂O₂) as a byproduct of electron transport. Rather than simply a toxic waste product, H₂O₂ functions as a second messenger, reversibly oxidizing cysteine residues (thiols) on regulatory proteins to switch their activity on and off. This redox-based signaling mechanism allows mitochondria to sense metabolic status and communicate it to the rest of the cell.
Additionally, the mitochondrial NAD+/NADH ratio serves as a master metabolic sensor. A low NAD+ level signals cellular stress and energy depletion, activating AMP-activated protein kinase (AMPK) and triggering a cascade of catabolic responses—energy-sparing pathways that downregulate biosynthesis and upregulate recycling of cellular components. This metabolic sensing function allows mitochondria to coordinate survival strategies during nutrient scarcity or stress, independent of energy depletion alone. See our coverage of New Studies for the latest research on metabolic sensing.
Mitochondrial DNA and Innate Immunity
Mitochondrial DNA (mtDNA) and mitochondrial RNA (mtRNA) released during mitochondrial stress or damage activate innate immune pathways, including the cGAS-STING pathway and Toll-like receptors. Recent immunological research shows that mtDNA acts as a damage-associated molecular pattern (DAMP), triggering interferon responses and inflammatory cytokine production when it leaks into the cytoplasm or is released into the bloodstream.
This function explains why mitochondrial damage in one tissue can trigger systemic inflammation. It also suggests that strategies targeting mitochondrial integrity may modulate immune function—a principle being explored in sepsis, autoimmune diseases, and chronic inflammatory conditions. Explore health rights and immunity topics for public-facing immunology content.
TCA Metabolites and Chromatin Remodeling
Metabolites produced by the tricarboxylic acid (TCA) cycle—citrate, α-ketoglutarate, succinate, and fumarate—are not merely intermediate steps toward energy production. These compounds serve as cofactors for epigenetic enzymes that remodel chromatin and determine which genes are expressed. Citrate is an acetyl-CoA precursor used by histone acetyltransferases; α-ketoglutarate is required by ten-eleven translocation (TET) enzymes and histone demethylases; succinate and fumarate inhibit these same enzymes, creating a metabolite-driven code that links metabolism directly to gene expression.
Epigenetic research demonstrates that shifting the TCA cycle flux changes which genes are activated or silenced, allowing mitochondrial metabolism to program cell identity and fate. This mechanism means that mitochondrial disease—or even temporary shifts in metabolic state—can directly alter the transcriptional landscape of a cell without changes to DNA sequence.
Mitochondria function as a central signaling hub coordinating at least five independent pathways: apoptosis via cytochrome c release, redox signaling through H₂O₂ and NAD+/NADH ratios, metabolic sensing via AMPK activation, innate immune activation through mtDNA/mtRNA release, and epigenetic programming via TCA cycle metabolites—collectively orchestrating cell survival, death, immunity, and gene expression far beyond their classical role in ATP synthesis.
— Cellular and molecular biology consensus literature
What this means
Frequently asked questions
Are mitochondrial diseases caused by energy failure alone?
No. While ATP depletion contributes to symptoms in some cases, many mitochondrial diseases reflect failure of signaling pathways—impaired apoptosis (allowing cancer development), redox dysregulation (causing oxidative stress), or epigenetic dysfunction (disrupting gene expression). Therapies targeting only energy production may miss critical disease mechanisms.
Can targeting mitochondrial function treat diseases beyond mitochondrial cytopathies?
Possibly. Since mitochondrial signaling coordinates apoptosis, immunity, and metabolism, modulating mitochondrial function may benefit cancer (improving cytochrome c release), autoimmune disease (reducing mtDNA-driven inflammation), and metabolic disease (enhancing NAD+-dependent AMPK pathways). Early-stage research is exploring this hypothesis.
What are practical ways to support mitochondrial health?
Regular aerobic exercise, fasting or caloric restriction (activating AMPK), adequate sleep, antioxidant-rich diet, and avoiding mitochondrial toxins (certain medications, heavy metals) support mitochondrial function. Emerging pharmaceuticals targeting NAD+ metabolism (NAD+ precursors, sirtuins) and mitochondrial dynamics are under development and clinical evaluation.
The emerging picture of mitochondrial biology reflects a shift from viewing these organelles as simple ATP factories to recognizing them as central command centers orchestrating multiple aspects of cellular and organismal physiology. This expanded framework opens new therapeutic avenues for diseases once thought unrelated to mitochondrial function and may explain why mitochondrial health is often preserved through the same interventions—exercise, metabolic health, sleep quality—that promote general wellness across disease and age groups.
Source: Mitochondria coordinate far more than ATP
Was this article helpful?
Disclaimer. This article is health journalism intended for general information and education. It is not medical advice and is not a substitute for professional diagnosis or treatment. Always consult a qualified healthcare provider about your individual circumstances. Full disclaimer →
Related Coverage
Editorial standards. This article was produced under the GMJ News editorial process, with oversight by the GMJ Editorial Board. Our editorial process. Spotted an error? Contact the editorial team.